Multivalent RNA motifs control alternative splicing

RNA-binding proteins (RBPs) regulate splicing according to position-dependent principles, which can be exploited for analysis of regulatory motifs. In collaboration with the laboratory of Prof Jernej Ule, we developed RNAmotifs, a method that evaluates the sequence around differentially regulated alternative exons to identify clusters of short and degenerate sequences, referred to as multivalent RNA motifs. We showed that diverse RBPs share basic positional principles, but differ in their propensity to enhance or repress exon inclusion. We now are interested in understanding how these principles work in cancer tissues and developing novel methods to disentangle such complexity.
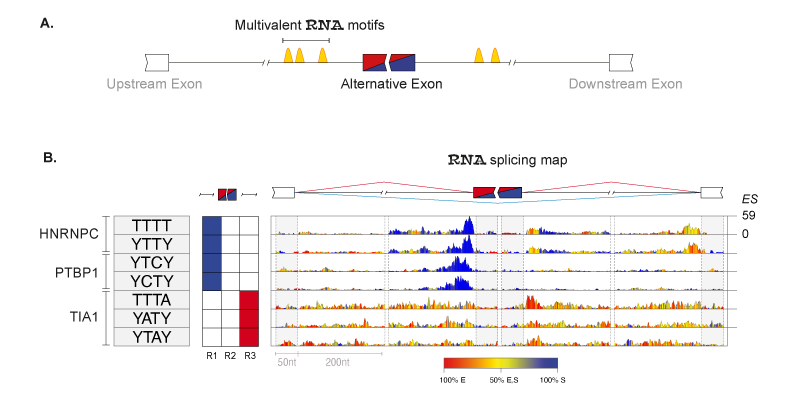
Principles of RNAmotifs algorithm
Principles of RNAmotifs algorithm. (A) Schematic representation of multivalent motifs surrounding the alternative spliced exon. (B) RNA splicing maps of multivalent RNA motifs enriched in the 'mixed' set of exons regulated by hnRNP C, PTBP1 and TIA.

We have now extended this framework into RNAmotifs 2.0. While the original method discovers enriched multivalent RNA motifs (MRMs) from a set of regulated exons, its key novelty is the association of MRMs with the RNA-binding proteins (RBPs) that recognise them. RNAmotifs 2.0 first calibrates, once, the positional binding principles of each RBP from orthogonal experimental evidence — eCLIP-seq binding profiles and knockdown RNA-seq — and optimises the RNAmotifs parameters per RBP. It then runs motif discovery on any new alternative-splicing dataset and matches the enriched MRMs to these RBP binding principles, producing a quantitative MRM–RBP association map that links each motif set to its candidate regulators, their RNA splicing maps and their enrichment across the exon regions. This turns motif discovery into a direct, testable readout of which RBPs drive a given splicing program.
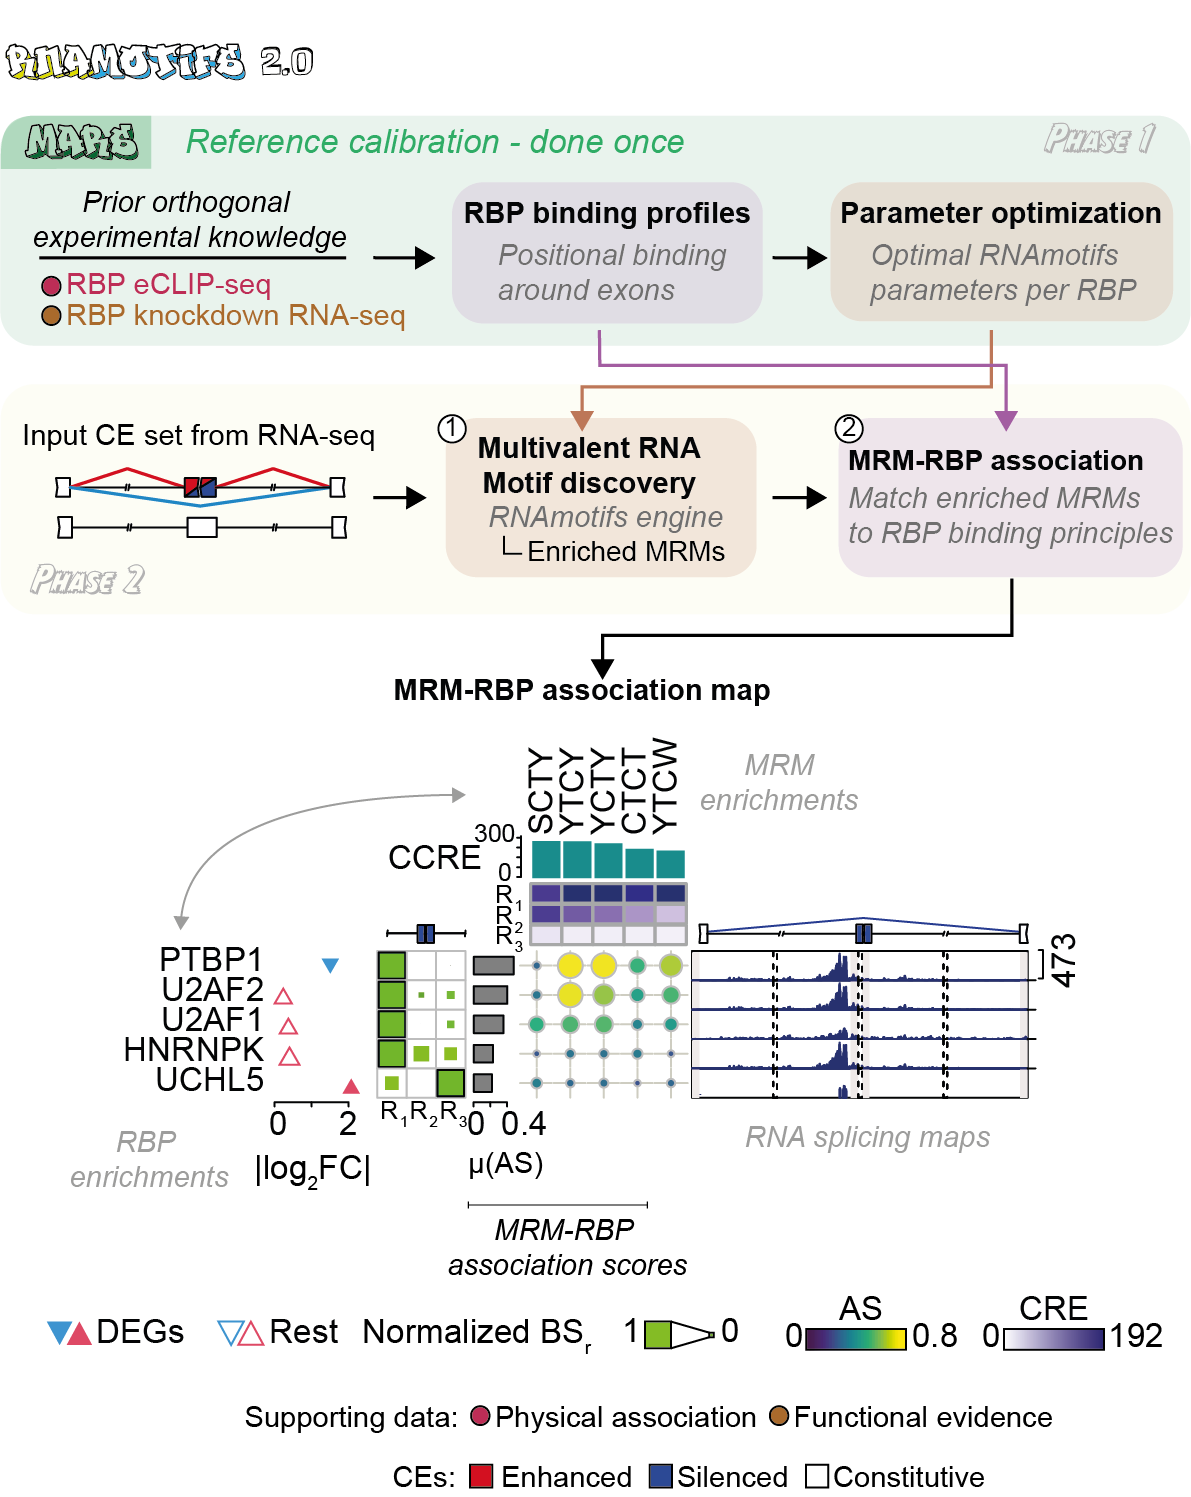
RNAmotifs 2.0 workflow and MRM–RBP association map
RNAmotifs 2.0 workflow. Phase 1 (reference calibration, done once): prior orthogonal experimental knowledge — RBP eCLIP-seq and RBP knockdown RNA-seq — is used to build positional RBP binding profiles around exons and to optimise the RNAmotifs parameters per RBP. Phase 2 (application): from an input set of cassette exons (CEs) from RNA-seq, (1) the RNAmotifs engine discovers enriched multivalent RNA motifs (MRMs), and (2) the enriched MRMs are matched to the calibrated RBP binding principles. The result is an MRM–RBP association map that integrates, per candidate RBP, its expression change (|log2FC|), the normalised binding scores across exon regions (R1, R2, R3), MRM enrichments, MRM–RBP association scores and RNA splicing maps, distinguishing enhanced, silenced and constitutive exons.

Details on RNAmotifs are here on Genome Biology, and on RNAmotifs 2.0 here on BMC Biology!


The transcriptional regulation of splicing factors

Alterations of protein-RNA interaction can lead to a variety of diseases, including cancer. The aberrant expression of RBPs has proven to have an oncogenic effect. Recently, a few studies have accessed AS deregulation as a general step in cancer onset showing that the upregulation of MYC leads to the abnormal expression of splicing factors that ultimately induces cell proliferation in brain tumours. We are interested in understanding the transcriptional regulation of splicing factors in cancer.

Our preliminary results in prostate cancer are available here on biorxiv!
Mutation load is not enough

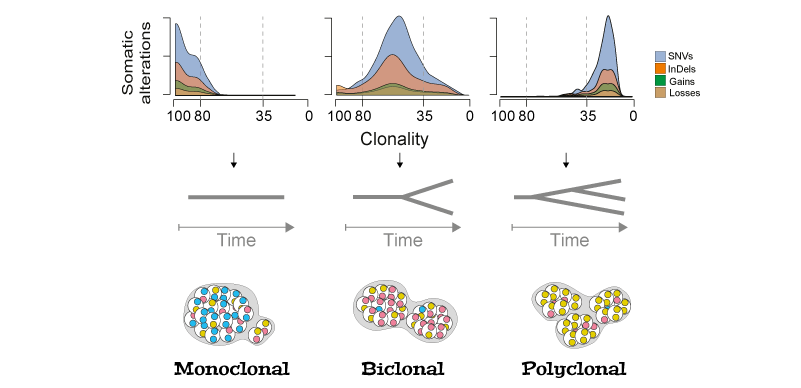
Cancer is a disease dominated by heterogeneity whose effects are evident in the evolution and treatment of the disease. In the laboratory of Prof Francesca Ciccarelli, we characterized the genetics of synchronous colorectal cancers and demonstrated that tumors of the same patient are genetically heterogeneous with clear therapeutic implications. To expand the number of "actionable" targets, relying only on the mutational load of tumors is not sufficient. We are currently studying somatic splicing aberrations to uncover novel potential therapeutic targets.

Tumor evolution reconstruction
Tumor evolution reconstruction. Schematic representation of tumor clonal evolution reconstruction. The density distributions of clonality for each type of alteration recapitulate the tumour clone composition.

Details on clonal evolution of synchronous colorectal cancers are here on Nature Communications!

We are currently leading the genomics analyses of Genomic Profiles Analysis in Children, Adolescents and Young Adult With Sarcomas (NCT04621201) trials.

"Less is more" in RNA-seq data analysis

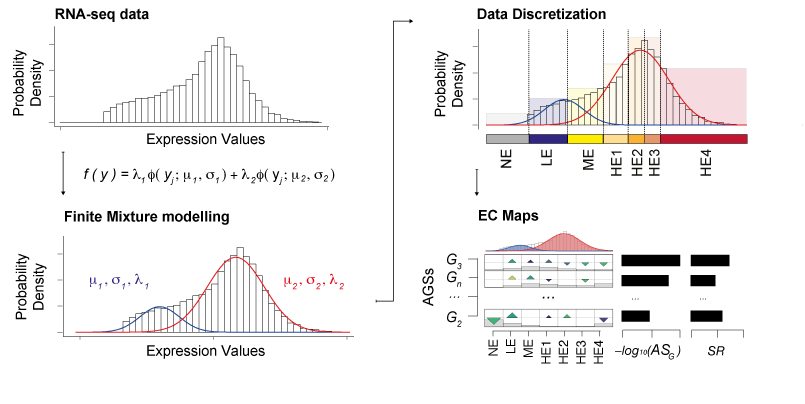
With the growth of high-throughput sequencing projects modern biology is facing novel bottlenecks due to Big Data issues. We addressed this challenge and introduced the novel concept of discretization of gene expression levels, which we derived from probabilistic modelling and shaped upon knowledge of RNA biology. Our Gene Set Enrichment Class Analysis (GSECA) algorithm exploits the bimodal behavior of RNA-sequencing gene expression profiles to identify altered gene sets in heterogeneous patient cohorts.

Schematic representation of GSECA algorithm
Schematic representation of GSECA algorithm. The algorithm proceeds through three sequential steps: (i) the sample-specific finite mixture modelling of gene expression distribution; (ii) the sample-specific discretization of expression values; and (iii) the statistical identification of altered gene sets (AGSs).

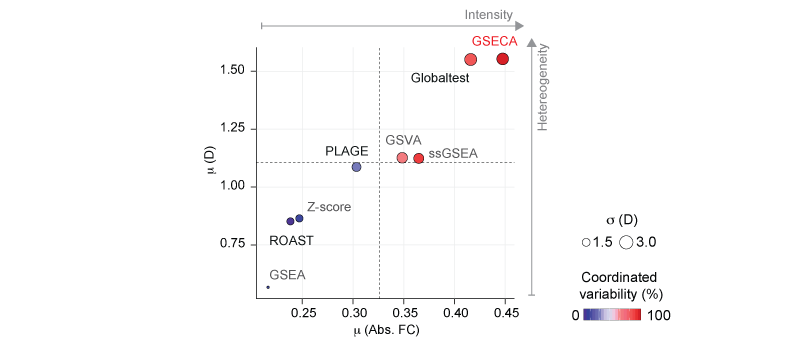
We showed that GSECA outperformed 'state-of-art' algorithms in handling gene sets characterized by heterogeneous expression changes. By boosting signal-to-noise ratio, GSECA can successfully manage the heterogeneity of thousands of samples.

Performance evaluation of gene set analysis algorithms
Performance evaluation of gene set analysis algorithms. Scatter plot of the mean absolute FC and dispersion averaged on the gene sets detected by each method.

With this work we introduced the paradigm shift of "less is more" in treating large heterogeneous RNA-seq datasets showing that it improves the detection of the altered biological processes in the phenotype of interest.

GSECA metaphor
GSECA metaphor. Heterogeneity can reveal a message if we look at it from a distance!

GSECA is out on Nucleic Acid Research, for any detail have a look there!


PTEN loss impacts on immune-related processes across cancers

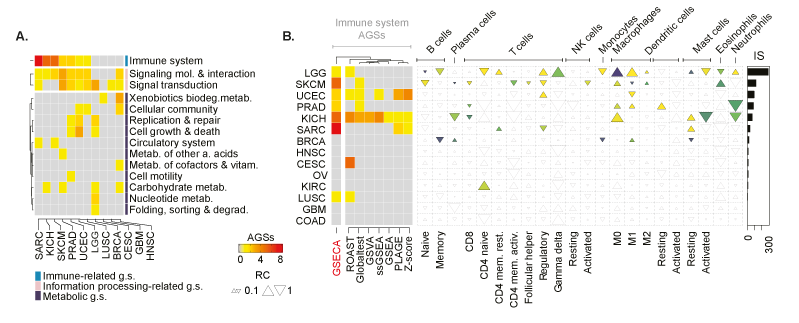
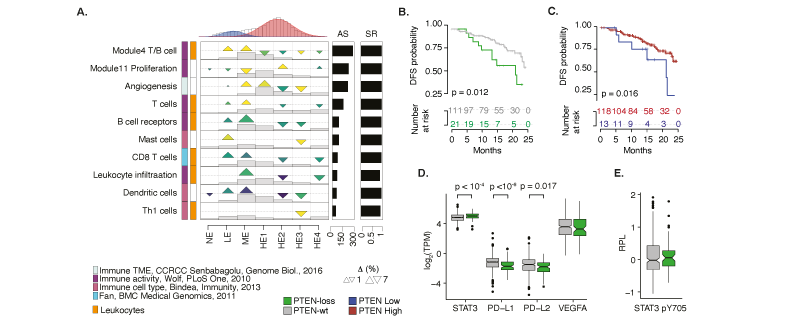
We generated a comprehensive assessment of the effect of PTEN loss across different cancer types. GSECA correctly highlighted the role of PTEN in controlling immune-related processes in the majority of cancer types, particularly in those showing a significant alteration of the tumor immune-microenvironment (TIME) composition.

Pan-cancer analysis of PTEN somatic loss
Pan-cancer analysis of PTEN somatic loss. (A) Heatmap showing the altered classes of gene sets across cancer types. (B) Heatmap shows the number of immune-related gene sets altered upon the loss of PTEN across cancer types.

Our analyses supported the notion that PTEN loss prostate cancers are non-T cell inflamed, or "cold", tumors.

Impact of PTEN loss in prostate cancer
Impact of PTEN loss in prostate cancer. (A) GSECA EC map showing altered immune expression signatures. (B) Disease-free survival curves. (C) DFS stratified on optimal PTEN expression level.

Details on our pancancer analysis of PTEN loss are here on Nucleic Acid Research!


Artificial Intelligence in transcriptomic Big Data

Artificial intelligence is becoming a fundamental asset for healthcare and life science research. AI is the pivotal tool to exploit the information available in genomic Big Data and ultimately "deliver" a medicine of precision.

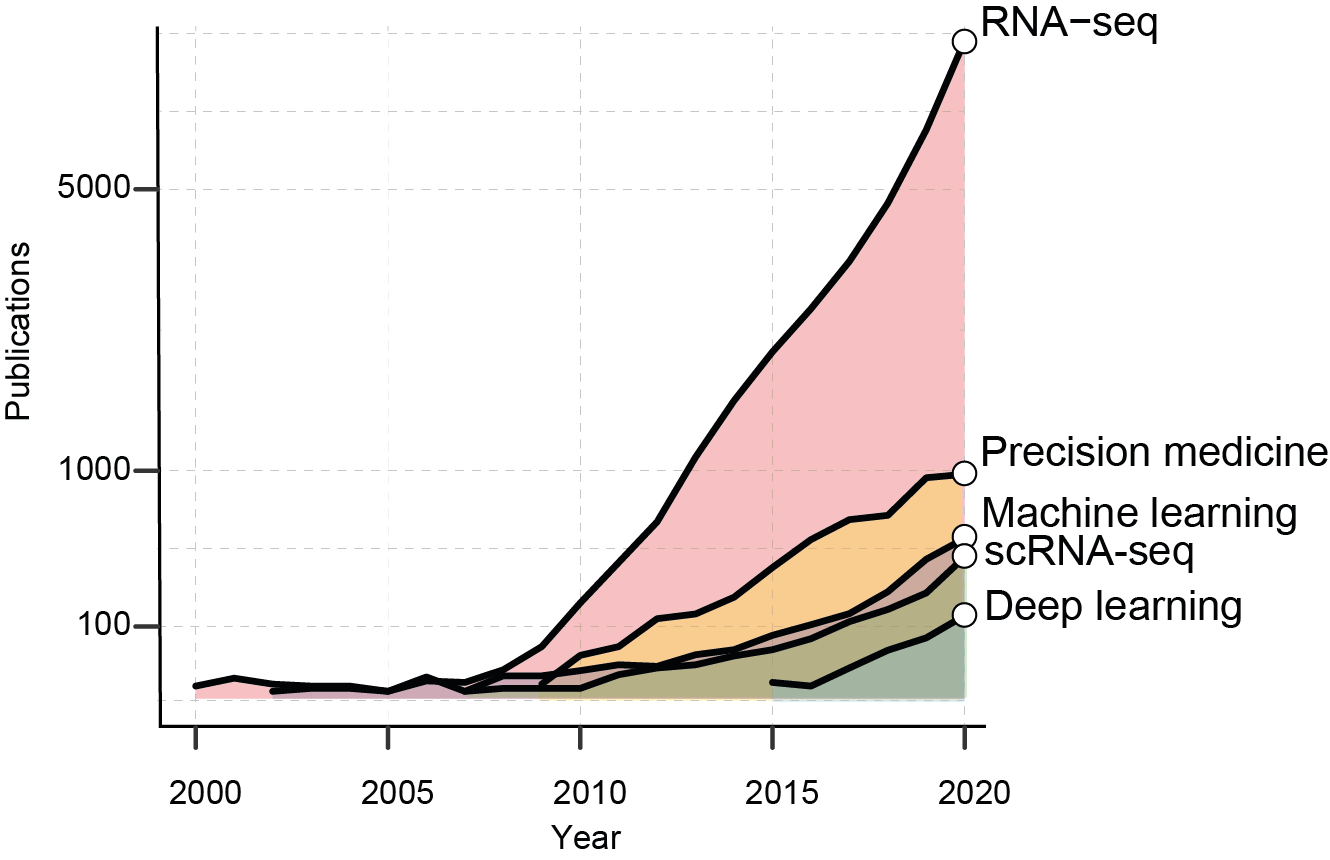
Growth of RNA-seq based studies
The growth of RNA-seq based studies. The graph shows the number of PubMed publications per year containing the reported keywords.

Machine and Deep Learning can leverage the heterogeneity of transcriptomic Big Data to achieve consistent predictions without the need of modeling the system of interest.

Artificial intelligence in cancer
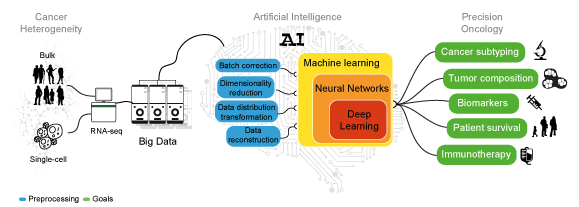
Artificial human intelligence. Sketch representing the analyses needed to decipher cancer heterogeneity and achieve an effective precision oncology.

Our review on Artificial Intelligence is out on International Journal of Molecular Sciences, for any detail have a look there! We also have an R-based tutorial for AI development that is available here.